Article Sections
Electronic scale properties of pristine stanene and tin forms using ab-initio methods
Loading author information...
Graphical Abstract & Abstract
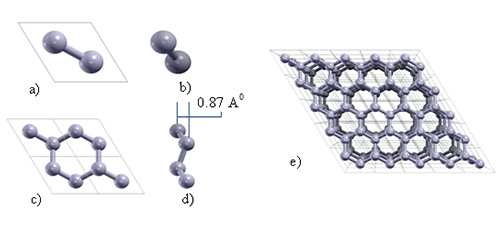
In the current study, elastic properties of stanene, a hexagonal honeycomb allotrope of tin is investigated using tools in computational material science. The simulations are performed using Quantum Espresso, an open-source package suit used for conducting ab-initio density functional theory simulations. The lattice structure of stanene analogous to the other group-IV elements 2D structures like graphene, silicene, and germanene. The relaxed structure of stanene in hexagonal honeycomb structure is found to have a lattice parameter . Unlike pristine graphene which has no buckling, stanene structure shows a buckling . All the calculations are carried out using generalized gradient approximation (GGA), and the exchange-correlation is treated using Perdew-Burke-Ernzerhof (PBE) functional. The cohesive energy of the structure is found to be . The calculations are conducted at groundstate without the inclusion of spin orbit coupling. The band structure, total and partial density of states a
Keywords
How to Cite
(2019). Electronic scale properties of pristine stanene and tin forms using ab-initio methods. Advanced Materials Letters, 10(1), 74-78. https://doi.org/10.5185/amlett.2019.2154